 |
OpenMS
|
Annotates PSMs with rescoring features derived from PeptDeep predictions. More...
#include <OpenMS/ANALYSIS/ID/PeptDeepRescoring.h>
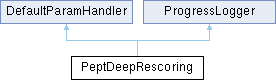
Classes | |
| struct | PSMRows |
| struct | Row |
| Row of the flat PSM work list built by annotate(). More... | |
Public Member Functions | |
| PeptDeepRescoring () | |
| Constructor. Sets the default parameters; the model paths start empty and must be set. | |
| ~PeptDeepRescoring () override | |
| Destructor. | |
| void | annotate (const PeakMap &spectra, std::vector< ProteinIdentification > &protein_ids, PeptideIdentificationList &peptide_ids) |
Annotates peptide_ids in place and registers the features for rescoring. | |
| void | selectNCE (const PeakMap &spectra, std::vector< ProteinIdentification > &protein_ids, const PeptideIdentificationList &peptide_ids) |
Selects the normalised collision energy of each run of peptide_ids as annotate() does, without predicting the spectra of all PSMs or adding features. | |
| void | selectNCE (const PeakMap &spectra, std::vector< ProteinIdentification > &protein_ids, const PeptideIdentificationList &peptide_ids, PeptDeepMS2Inference &ms2_inference) |
| double | getUsedNCE () const |
| double | getRTCalibrationError () const |
| Size | getNumPSMs () const |
| Number of PSMs (peptide hits) seen by the last annotate() call. | |
| Size | getNumPredictedPSMs () const |
 Public Member Functions inherited from DefaultParamHandler | |
| DefaultParamHandler (const std::string &name) | |
| Constructor with name that is displayed in error messages. | |
| DefaultParamHandler (const DefaultParamHandler &rhs) | |
| Copy constructor. | |
| virtual | ~DefaultParamHandler () |
| Destructor. | |
| DefaultParamHandler & | operator= (const DefaultParamHandler &rhs) |
| Assignment operator. | |
| virtual bool | operator== (const DefaultParamHandler &rhs) const |
| Equality operator. | |
| void | setParameters (const Param ¶m) |
| Sets the parameters. | |
| const Param & | getParameters () const |
| Non-mutable access to the parameters. | |
| const Param & | getDefaults () const |
| Non-mutable access to the default parameters. | |
| const std::string & | getName () const |
| Non-mutable access to the name. | |
| void | setName (const std::string &name) |
| Mutable access to the name. | |
| const std::vector< std::string > & | getSubsections () const |
| Non-mutable access to the registered subsections. | |
| Public Member Functions inherited from ProgressLogger | |
| ProgressLogger () | |
| Constructor. | |
| virtual | ~ProgressLogger () |
| Destructor. | |
| ProgressLogger (const ProgressLogger &other) | |
| Copy constructor. | |
| ProgressLogger & | operator= (const ProgressLogger &other) |
| Assignment Operator. | |
| void | setLogType (LogType type) const |
| Sets the progress log that should be used. The default type is NONE! | |
| LogType | getLogType () const |
| Returns the type of progress log being used. | |
| void | setLogger (ProgressLoggerImpl *logger) |
| Sets the logger to be used for progress logging. | |
| void | startProgress (SignedSize begin, SignedSize end, const std::string &label) const |
| Initializes the progress display. | |
| void | setProgress (SignedSize value) const |
| Sets the current progress. | |
| void | endProgress (UInt64 bytes_processed=0) const |
| void | nextProgress () const |
| increment progress by 1 (according to range begin-end) | |
Static Public Member Functions | |
| static StringList | featureNames () |
| Names of the features this class adds to each PSM, in a stable order. | |
| Static Public Member Functions inherited from DefaultParamHandler | |
| static void | writeParametersToMetaValues (const Param &write_this, MetaInfoInterface &write_here, const std::string &key_prefix="") |
| Writes all parameters to meta values. | |
Static Public Attributes | |
| static constexpr const char * | NCE_META_VALUE = "peptdeep_nce" |
Protected Member Functions | |
| void | updateMembers_ () override |
| This method is used to update extra member variables at the end of the setParameters() method. | |
| bool | startCall_ (const PeptideIdentificationList &peptide_ids, const StringList &models, const std::string &what) |
| void | selectNCE_ (const PeakMap &spectra, std::vector< ProteinIdentification > &protein_ids, const PeptideIdentificationList &peptide_ids, PeptDeepMS2Inference &ms2_inference) |
selectNCE() with the session ms2_inference | |
| std::vector< Size > | confidentRows_ (const std::vector< Row > &rows, const std::vector< Size > &run_rows, bool higher_better) const |
| double | selectRunNCE_ (const std::map< std::string, double > &collision_energies, const PeptideIdentificationList &peptide_ids, const PSMRows &psms, const std::vector< Size > &run_rows, const std::vector< Size > &confident, PeptDeepMS2Inference &ms2_inference, double ¢re, double &median_cosine) |
| void | annotateRun_ (const std::string &run_id, const std::map< std::string, double > &collision_energies, PeptideIdentificationList &peptide_ids, const PSMRows &psms, const std::vector< Size > &run_rows, bool higher_better, PeptDeepMS2Inference &ms2_inference, PeptDeepRTInference &rt_inference) |
| Protected Member Functions inherited from DefaultParamHandler | |
| void | defaultsToParam_ () |
| Updates the parameters after the defaults have been set in the constructor. | |
Static Protected Member Functions | |
| static PSMRows | gatherPSMs_ (const PeptideIdentificationList &peptide_ids) |
| static std::map< std::string, double > | collisionEnergies_ (const PeakMap &spectra) |
| static void | storeNCE_ (std::vector< ProteinIdentification > &protein_ids, const std::map< std::string, double > &nce_by_run) |
Protected Attributes | |
| std::string | ms2_model_ |
| std::string | rt_model_ |
| double | nce_ {-1.0} |
| double | nce_grid_halfwidth_ {6.0} |
| double | nce_grid_step_ {2.0} |
| double | nce_fallback_ {30.0} |
| Size | nce_sample_size_ {2000} |
| std::string | instrument_ |
| double | calibration_quantile_ {0.5} |
| std::string | rt_model_type_ {"b_spline"} |
| Size | rt_num_nodes_ {5} |
| int | ms2_min_ordinal_ {1} |
| double | ms2_strong_fraction_ {0.05} |
| Size | batch_size_ {64} |
| Int | threads_ {4} |
| double | used_nce_ {-1.0} |
| double | rt_calibration_error_ {-1.0} |
| Size | num_psms_ {0} |
| Size | num_predicted_psms_ {0} |
| Protected Attributes inherited from DefaultParamHandler | |
| Param | param_ |
| Container for current parameters. | |
| Param | defaults_ |
| Container for default parameters. This member should be filled in the constructor of derived classes! | |
| std::vector< std::string > | subsections_ |
| Container for registered subsections. This member should be filled in the constructor of derived classes! | |
| std::string | error_name_ |
| Name that is displayed in error messages during the parameter checking. | |
| bool | check_defaults_ |
| If this member is set to false no checking if parameters in done;. | |
| bool | warn_empty_defaults_ |
| If this member is set to false no warning is emitted when defaults are empty;. | |
| Protected Attributes inherited from ProgressLogger | |
| LogType | type_ |
| time_t | last_invoke_ |
| ProgressLoggerImpl * | current_logger_ |
Additional Inherited Members | |
| Public Types inherited from ProgressLogger | |
| enum | LogType { CMD , GUI , NONE } |
| Possible log types. More... | |
| Static Protected Attributes inherited from ProgressLogger | |
| static std::atomic< int > | recursion_depth_ |
Annotates PSMs with rescoring features derived from PeptDeep predictions.
Compares every PSM against an AlphaPeptDeep prediction of the same peptide and stores the agreement as per-hit features, which are then registered in the search parameters' extra_features list so that PercolatorAdapter picks them up:
ms2_cosine, ms2_spectral_angle, ms2_pearson, ms2_frac_pred_found – predicted versus observed fragment intensitiesrt_abs_error – observed retention time versus a per-run calibration of the predicted retention timeObserved intensities are taken from the PSM's own peak annotations, so no second pass over the spectra is needed; the search engine must have been run with fragment annotation enabled (ProSE's annotate:PSM default of ALL does this). Predicted ions that were not matched contribute zero on the observed side, which is what makes a wrong peptide score badly rather than simply score on fewer ions.
Two per-run quantities are calibrated from the data rather than being asked of the user, because both are properties of the run and neither has a safe global default:
nce set to a positive value the scan is skipped and that value is used directly.Confident PSMs are selected by search score rather than by spectral similarity: using similarity here would tie the RT feature to the MS2 features and let one feature's errors propagate into the other. Only the best-scoring hit of each spectrum, and only if it is not a decoy, is a candidate, so the calibration does not depend on how many hits per spectrum the search reports: with ten, lower-ranked candidates and decoys would otherwise make up most of the better-scoring half.
Parameters are not pulled into the generated documentation with @htmlinclude: that requires the class to be registered in DefaultParamHandlerDocumenter, which cannot instantiate a type that only exists in ONNX-enabled builds. ProSEAlgorithm omits it for the same reason.
| struct OpenMS::PeptDeepRescoring::PSMRows |
The PSMs of annotate() and selectNCE(): a row per hit with a sequence, the distinct (sequence, charge) pairs to predict, and the rows of each run (by run identifier, in a stable order)
| Class Members | ||
|---|---|---|
| vector< Row > | rows | |
| map< string, vector< Size > > | rows_by_run | |
| vector< float > | uniq_charge | |
| vector< string > | uniq_seq | |
| struct OpenMS::PeptDeepRescoring::Row |
Row of the flat PSM work list built by annotate().
| Class Members | ||
|---|---|---|
| bool | calibration_candidate | best hit of its spectrum and not a decoy |
| Size | hit | index into that identification's hits |
| Size | key | index into the de-duplicated (sequence, charge) list |
| Size | len | peptide length |
| Size | pi | index into the PeptideIdentificationList |
| double | score | search score, used to pick the calibration set |
Constructor. Sets the default parameters; the model paths start empty and must be set.
|
override |
Destructor.
| void annotate | ( | const PeakMap & | spectra, |
| std::vector< ProteinIdentification > & | protein_ids, | ||
| PeptideIdentificationList & | peptide_ids | ||
| ) |
Annotates peptide_ids in place and registers the features for rescoring.
| [in] | spectra | Spectra the identifications were made on; supplies retention time and, for the NCE scan, the instrument's reported collision energy. |
| [in,out] | protein_ids | Search metadata; the feature names are appended to the extra_features of each run that has PSMs, and the NCE its predictions used is stored on the run as the double meta value NCE_META_VALUE (see below). |
| [in,out] | peptide_ids | PSMs to annotate. Every hit receives every feature, so that no run ends up with a feature present on only some of its PSMs. |
Multiple runs in one call are fine: PSMs are partitioned by PeptideIdentification::getIdentifier() and each run gets its own NCE scan and its own RT calibration.
Each annotated ProteinIdentification gets the meta value NCE_META_VALUE ("peptdeep_nce", a double): the NCE its run's predictions used – the one selected by the scan, or nce when that is set. It lets a consumer of the result predict spectra at the same energy the annotation used. A ProteinIdentification without PSMs is not annotated and gets none. If no run identifier matches any ProteinIdentification, the features go on the first one as before, and so does the value, but only when all runs used the same NCE.
out_merged and IDMergerAlgorithm, which remap every identifier onto a single merged run, or IDMerger with merge_proteins_add_PSMs (its default appends runs and is therefore safe). Both calibrated quantities genuinely differ between runs: six files of one HLA ligand atlas dataset (PXD019643) calibrated to median RT residuals between 98 s and 625 s. On already-collapsed input this silently fits one NCE and one RT curve across all of them, and nothing here can detect it after the fact – the merged input is indistinguishable from a genuine single run.| Exception::FileNotFound | if a configured model file does not exist. |
| Exception::MissingInformation | if no PSM carries peak annotations, since every MS2 feature would otherwise be silently zero. |
|
protected |
Annotates the PSMs of a single identification run. Each run gets its own collision energy and its own retention-time calibration, since neither transfers across runs with different gradients or acquisition settings.
|
staticprotected |
The collision energy of each MS2 spectrum of spectra by native ID (native IDs recorded with differing energies are left out, with a warning)
|
protected |
The confident rows of a run (run_rows of rows): its calibration candidates (all rows if it has none), best search score first, without the calibration_quantile held out
|
static |
Names of the features this class adds to each PSM, in a stable order.
|
staticprotected |
The PSMs of peptide_ids. Throws Exception::MissingInformation if no PSM carries peak annotations, and warns if only some runs do.
| Size getNumPredictedPSMs | ( | ) | const |
Number of PSMs of the last annotate() call that received a model prediction and carry peak annotations, i.e. whose MS2 features are computed rather than zero-filled. 0 if annotate() returned early or predicted nothing.
| Size getNumPSMs | ( | ) | const |
Number of PSMs (peptide hits) seen by the last annotate() call.
| double getRTCalibrationError | ( | ) | const |
Median absolute RT residual over the calibration set of the run annotated last. -1 if annotate() has not run, or could not calibrate.
| double getUsedNCE | ( | ) | const |
Normalised collision energy used for the run annotated last (after any scan). -1 if annotate() has not run, or returned before selecting one.
| void selectNCE | ( | const PeakMap & | spectra, |
| std::vector< ProteinIdentification > & | protein_ids, | ||
| const PeptideIdentificationList & | peptide_ids | ||
| ) |
Selects the normalised collision energy of each run of peptide_ids as annotate() does, without predicting the spectra of all PSMs or adding features.
For a caller that predicts spectra itself at the energy annotate() would use (e.g. ProSE's -qpx_extended for runs that were not rescored with PeptDeep): the same selection on the same sample of confident PSMs (the scan of the grid, or nce if that is set), and the NCE stored on the runs as NCE_META_VALUE, exactly as annotate() stores it. Needs only ms2_model. Logs no "[PeptDeepRescoring]" lines of its own besides warnings about the input (shared with annotate()); its progress is that of the scan ("Selecting collision energy...").
| [in] | spectra | As for annotate() |
| [in,out] | protein_ids | The runs; each run with PSMs gets NCE_META_VALUE |
| [in] | peptide_ids | The PSMs, as for annotate(); not changed |
| Exception::FileNotFound | if ms2_model does not exist. |
| Exception::MissingInformation | if ms2_model is not set, or no PSM carries peak annotations. |
| void selectNCE | ( | const PeakMap & | spectra, |
| std::vector< ProteinIdentification > & | protein_ids, | ||
| const PeptideIdentificationList & | peptide_ids, | ||
| PeptDeepMS2Inference & | ms2_inference | ||
| ) |
selectNCE() with the session of the MS2 model ms2_inference (e.g. one for all inputs of a run) instead of one of ms2_model; its threads and batch size apply
|
protected |
selectNCE() with the session ms2_inference
|
protected |
The NCE of a run (run_rows of psms): 'nce' if set, else the grid scan on a sample of its confident rows. With the scan, centre is the centre of its grid and median_cosine the median cosine of the selected NCE (else both unchanged).
|
protected |
The start of annotate() and selectNCE(): resets the diagnostics; throws Exception::MissingInformation (what) if one of models is not set, Exception::FileNotFound if one does not exist; false if there are no PSMs
|
staticprotected |
Stores the NCE of each run of nce_by_run on the run of protein_ids with its identifier (NCE_META_VALUE); if none has one of these identifiers, on the first run, provided that all NCEs agree
|
overrideprotectedvirtual |
This method is used to update extra member variables at the end of the setParameters() method.
Also call it at the end of the derived classes' copy constructor and assignment operator.
The default implementation is empty.
Reimplemented from DefaultParamHandler.
|
protected |
|
protected |
|
protected |
|
protected |
|
protected |
|
protected |
|
protected |
|
protected |
|
protected |
|
protected |
|
staticconstexpr |
Name of the run meta value ("peptdeep_nce", a double) in which annotate() stores, on each annotated ProteinIdentification, the normalised collision energy its predictions used. It is written to idXML with the run, so consumers of stored results can rely on the name.
|
protected |
|
protected |
|
protected |
|
protected |
|
protected |
|
protected |
|
protected |
|
protected |
|
protected |
 1.9.8
